New developments in medical research offer a significant ray of hope for individuals afflicted with Autosomal Dominant Optic Atrophy (ADOA), a prevalent inherited condition leading to visual impairment. Recent scientific endeavors have pinpointed a crucial therapeutic avenue: the deactivation of the SARM1 protein. This innovative approach has shown considerable promise in safeguarding retinal ganglion cells and maintaining visual function in laboratory studies, potentially revolutionizing future treatments for this progressive disorder.
Autosomal Dominant Optic Atrophy (ADOA) represents the most common inherited optic neuropathy, often manifesting subtly during childhood. Initial symptoms may include blurred vision, difficulty with reading, or merely failing an eye examination. However, these seemingly minor indicators mask a relentless progression towards irreversible vision loss affecting both eyes. This deterioration is primarily caused by the degeneration of retinal ganglion cells (RGCs), the neural pathways crucial for transmitting visual information from the eyes to the brain. In most instances, this damage is attributed to mutations within the OPA1 gene, which disrupts mitochondrial activity—essential for cellular energy production and health.
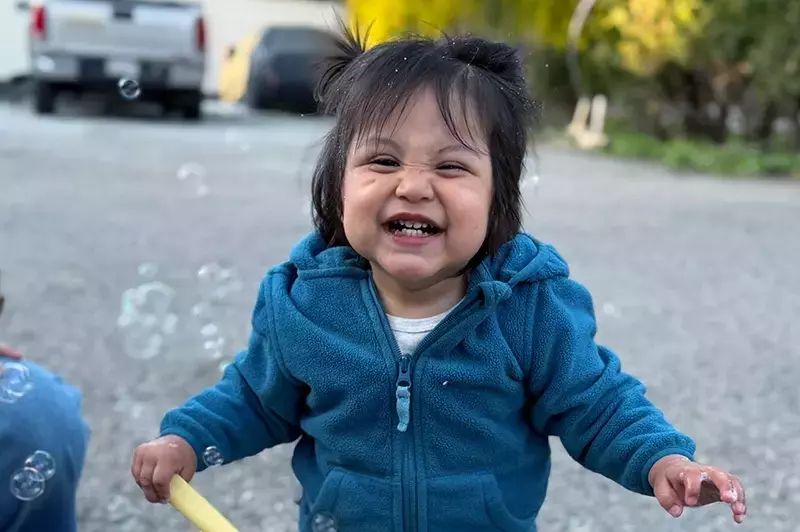
A groundbreaking study led by Dr. Thomas Schwarz and Dr. Chen Ding at Boston Children’s Hospital’s Schwarz Lab has uncovered a highly promising therapeutic target to counteract the damage to RGCs. Intriguingly, ADOA was not initially a primary focus of their research until 2019. The impetus for this pivotal shift came from a family whose daughter carries the genetic predisposition for the disease. They approached the lab with a heartfelt plea: to explore whether the lab's ongoing research into mitochondrial dysfunction could be adapted to address ADOA and, crucially, preserve their daughter’s sight. This poignant request brought a deeply personal dimension to the team's scientific pursuits, with the family’s unwavering support playing a vital role in transforming a nascent concept into a viable treatment strategy.
Dr. Schwarz highlighted the profound impact of this encounter, stating, \"It compelled us to collaborate and consider what impactful work we could undertake to help.\" The Schwarz Lab's findings, recently published in the Journal of Clinical Investigation, demonstrate that either the deletion or inactivation of a single protein, SARM1, can effectively preserve RGCs and sustain vision in mouse models. SARM1 is well-known for its role in initiating axon degeneration—the process where the signal-transmitting portion of a nerve cell breaks down. In the context of ADOA, the activation of SARM1 is what directly contributes to the damage observed in retinal ganglion cells.
Dr. Ding elaborated on the significance of their discovery, noting, \"Most treatments for neurodegenerative conditions like ADOA aim to prolong cell life or decelerate damage progression. What we've uncovered is essentially a molecular 'off-switch' for the very process that instigates cell death.\" When researchers removed SARM1 from mice harboring the OPA1 mutation, the RGCs retained their functionality, and the mice maintained their visual acuity. Dr. Schwarz emphasized the magnitude of this breakthrough, stating, \"In a field where effective interventions have been remarkably scarce, this represents a monumental stride.\"
With crucial backing from the affected girl’s family and a grant from the Advancium Health Network, a charitable organization dedicated to advancing therapies for rare diseases, the Schwarz Lab is now exploring methods to inhibit SARM1 pharmacologically, moving beyond genetic manipulation. Dr. Ding explained, \"If the SARM1 inhibition strategies we are currently testing in the lab prove effective, the subsequent phase will involve assessing if inhibiting it in human patients can yield similar nerve-protective outcomes.\" To achieve this, the research team plans to evaluate ASHA-624, a novel pharmaceutical agent designed to block SARM1 by stabilizing the protein in an inactive conformation, thereby preventing it from initiating axonal degeneration.
For healthcare professionals managing children with ADOA and for their families, Dr. Schwarz and Dr. Ding believe this is a truly transformative period. As SARM1-targeted therapies advance towards clinical trials, they underscore the critical importance of early genetic diagnosis. Such early identification will be paramount in pinpointing patients who could most benefit from future treatments, offering them a genuine prospect of preserving their vision rather than merely slowing its decline. Dr. Schwarz articulated the overarching sentiment: \"SARM1 inhibition offers a novel paradigm for understanding ADOA. We are optimistic that a viable therapy is within our grasp, and we are committed to accelerating its development with utmost urgency.\"